Unit 3
Chemical Kinetics
Objectives
After studying this Unit, you will be able to
- define the average and instantaneous rate of a reaction; - express the rate of a reaction in terms of change in concentration of either of the reactants or products with time; - distinguish between elementary and complex reactions; - differentiate between the molecularity and order of a reaction; - define rate constant; - discuss the dependence of rate of reactions on concentration, temperature and catalyst; - derive integrated rate equations for the zero and first order reactions; - determine the rate constants for zeroth and first order reactions; - describe collision theory.
The discipline of Chemical Kinetics offers an understanding of the processes underlying chemical transformations.
The fundamental essence of chemistry lies in the investigation of change. Through chemical reactions, substances with clearly defined properties are transmuted into distinct substances possessing different characteristics. In the context of any chemical reaction, chemists typically seek to determine:
(a) the practicability of a chemical reaction, which is ascertainable through thermodynamics (given that a reaction is considered feasible when $\Delta G < 0$ at constant temperature and pressure); (b) the degree of advancement a reaction will exhibit, which can be established using chemical equilibrium principles; (c) the velocity of a reaction, signifying the duration required for it to achieve equilibrium.
Beyond merely understanding the feasibility and extent of a chemical process, comprehending its reaction rate and the variables governing it is crucial for a comprehensive grasp of the phenomenon. Consider, for instance, the factors that dictate the speed of food spoilage. Or how might one formulate a quick-setting substance suitable for dental restorations? Furthermore, what mechanisms regulate the combustion velocity of fuel within an automotive engine? These inquiries are addressed by the domain of chemistry known as chemical kinetics, which is dedicated to investigating reaction rates and their underlying mechanisms. The term 'kinetics' itself originates from the Greek 'kinesis', signifying motion. While thermodynamics solely informs us about a reaction's potential for occurrence, chemical kinetics elucidates its actual pace. For instance, although thermodynamic evidence suggests the transformation of diamond into graphite, the actual rate of this conversion is exceedingly sluggish, rendering the alteration imperceptible over human timescales. Consequently, a common perception holds that diamond possesses eternal stability.
Investigations in kinetics not only enable the quantification of a chemical reaction's velocity but also elucidate the environmental parameters capable of modifying these reaction rates. Variables including reactant concentration, ambient temperature, applied pressure, and the presence of a catalyst all influence the reaction's speed. From a macroscopic perspective, the primary focus is on the quantities of substances consumed or generated, and the respective rates of their disappearance or appearance. Conversely, at the molecular scale, discussions revolve around the reaction mechanisms, particularly the orientation and energetic state of molecules during collisional events.
Within this Unit, the concepts of average and instantaneous reaction rates, along with their influencing factors, will be examined. Additionally, foundational principles pertaining to the collision theory of reaction rates will be introduced. Nonetheless, to fully grasp these topics, it is imperative to first establish an understanding of the reaction rate itself.
3.1 Rate of a Chemical Reaction
The pace at which chemical transformations unfold exhibits considerable variability. For instance, certain processes, such as ionic reactions, demonstrate extreme rapidity; a classic illustration is the instantaneous precipitation of silver chloride upon mixing aqueous solutions of silver nitrate and sodium chloride. Conversely, some reactions are notably slow, exemplified by the gradual rusting of iron when exposed to air and moisture. Furthermore, there exist reactions, like the inversion of cane sugar and the hydrolysis of starch, that proceed at a moderate velocity. Can you identify additional instances for each of these reaction rate categories?
As you are likely aware, the velocity of an automobile is quantified by the alteration in its position or the distance covered within a specific time interval. Analogously, the velocity of a chemical reaction, or its rate, is defined as the change in the concentration of either a reactant or a product per unit of time. More precisely, this rate can be articulated in two principal ways:
(i) the rate at which the concentration of any reactant diminishes, or (ii) the rate at which the concentration of any product increases.
Let us consider a hypothetical reaction, assuming that the system's volume remains constant:
$R \rightarrow P$
Here, one mole of reactant R converts into one mole of product P. If $\left[\mathrm{R}\right]{1}$ and $\left[\mathrm{P}\right]{1}$ denote the concentrations of R and P, respectively, at time $t_{1}$, and $\left[\mathrm{R}\right]{2}$ and $\left[\mathrm{P}\right]{2}$ represent their concentrations at a later time $t_{2}$, then the changes can be expressed as:
$\begin{array}{l} \Delta t = t_{2} - t_{1} \ \Delta [\mathrm{R}] = [\mathrm{R}]{2} - [\mathrm{R}]{1} \ \Delta [\mathrm{P}] = [\mathrm{P}]{2} - [\mathrm{P}]{1} \ \end{array}$
In the expressions above, the square brackets signify molar concentration.
The rate at which R disappears is given by:
$= \frac{\text{Decrease in concentration of R}}{\text{Time taken}} = - \frac{\Delta [\mathrm{R}]}{\Delta t} \tag{3.1}$
The rate at which P appears is given by:
$= \frac {\text {Increase in concentration of P}}{\text {Time taken}} = + \frac {\Delta [ \mathrm {P} ]}{\Delta t} \tag {3.2}$
Since $\Delta[\mathrm{R}]$ inherently yields a negative value (due to the diminishing concentration of reactants), it is multiplied by $-1$ to ensure that the reaction rate is reported as a positive quantity.
Equations (3.1) and (3.2), as presented above, characterize the average reaction rate, denoted as $r_{av}$.
The average rate is contingent upon the observed change in the concentrations of reactants or products, along with the duration over which this change occurs (Fig. 3.1).
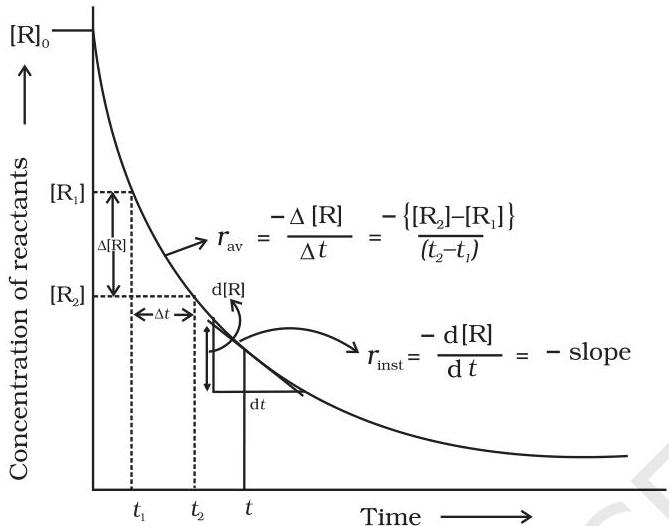
Fig. 3.1: Instantaneous and average rate of a reaction
Units of rate of a reaction
Based on equations (3.1) and (3.2), it can be established that the standard units for reaction rate are expressed as concentration per unit time (concentration time$^{-1}$). For instance, if concentration is quantified in moles per liter (mol L$^{-1}$) and time in seconds, the resulting units will be mol L$^{-1}$s$^{-1}$. Nevertheless, in the context of gaseous reactions, where gas concentrations are typically indicated by their partial pressures, the units for the rate equation will consequently be atm s$^{-1}$.
Given the concentrations of $\mathrm{C_4H_9Cl}$ (butyl chloride) recorded at various time points below, proceed to calculate the average reaction rate:
$\mathrm {C} _ {4} \mathrm {H} _ {9} \mathrm {C l} + \mathrm {H} _ {2} \mathrm {O} \rightarrow \mathrm {C} _ {4} \mathrm {H} _ {9} \mathrm {O H} + \mathrm {H C l}$
across the specified time intervals.
| t/s | 0 | 50 | 100 | 150 | 200 | 300 | 400 | 700 | 800 |
|---|---|---|---|---|---|---|---|---|---|
| [C4H9Cl]/mol L$^{-1}$ | 0.100 | 0.0905 | 0.0820 | 0.0741 | 0.0671 | 0.0549 | 0.0439 | 0.0210 | 0.017 |
The average rate can be ascertained by determining the change in concentration over distinct time intervals and subsequently dividing $\Delta[\mathrm{R}]$ by $\Delta t$, as illustrated in Table 3.1.
Table 3.1: Average rates of hydrolysis of butyl chloride
| [C4H9Cl]t₁ (mol L−1) |
[C4H9Cl]t₂ (mol L−1) |
t1 (s) |
t2 (s) |
rav × 104 (mol L−1 s−1) |
|---|---|---|---|---|
| 0.1000 | 0.0905 | 0 | 50 | 1.90 |
| 0.0905 | 0.0820 | 50 | 100 | 1.70 |
| 0.0820 | 0.0741 | 100 | 150 | 1.58 |
| 0.0741 | 0.0671 | 150 | 200 | 1.40 |
| 0.0671 | 0.0549 | 200 | 300 | 1.22 |
| 0.0549 | 0.0439 | 300 | 400 | 1.10 |
| 0.0439 | 0.0335 | 400 | 500 | 1.04 |
| 0.0210 | 0.0170 | 700 | 800 | 0.40 |
Note on Rate Calculation: rav = − ([C4H9Cl]t₂ − [C4H9Cl]t₁) / (t2 − t1)
As observed in Table 3.1, the average reaction rate diminishes from $1.90 \times 10^{-4} , \mathrm{mol} , \mathrm{L}^{-1} , \mathrm{s}^{-1}$ to $0.4 \times 10^{-4} , \mathrm{mol} , \mathrm{L}^{-1} , \mathrm{s}^{-1}$. However, the average rate is insufficient for predicting the reaction rate at a specific moment in time, as it remains constant throughout the calculated interval. Consequently, to quantify the rate at a precise instant, we determine the instantaneous rate. This is achieved by considering the average rate over an infinitesimally small time interval, denoted as $dt$ (i.e., as $\Delta t$ approaches zero). Thus, mathematically, for an infinitesimally small $dt$, the instantaneous rate is defined as:
$r_{\mathrm{av}} = \frac{-\Delta[\mathrm{R}]}{\Delta t} = \frac{\Delta[\mathrm{P}]}{\Delta t} \tag{3.3}$
$\mathrm{As} \quad \Delta t \rightarrow 0 \quad \text{or} \quad r_{\mathrm{inst}} = \frac{-\mathrm{d}[\mathrm{R}]}{\mathrm{d}t} = \frac{\mathrm{d}[\mathrm{P}]}{\mathrm{d}t}$
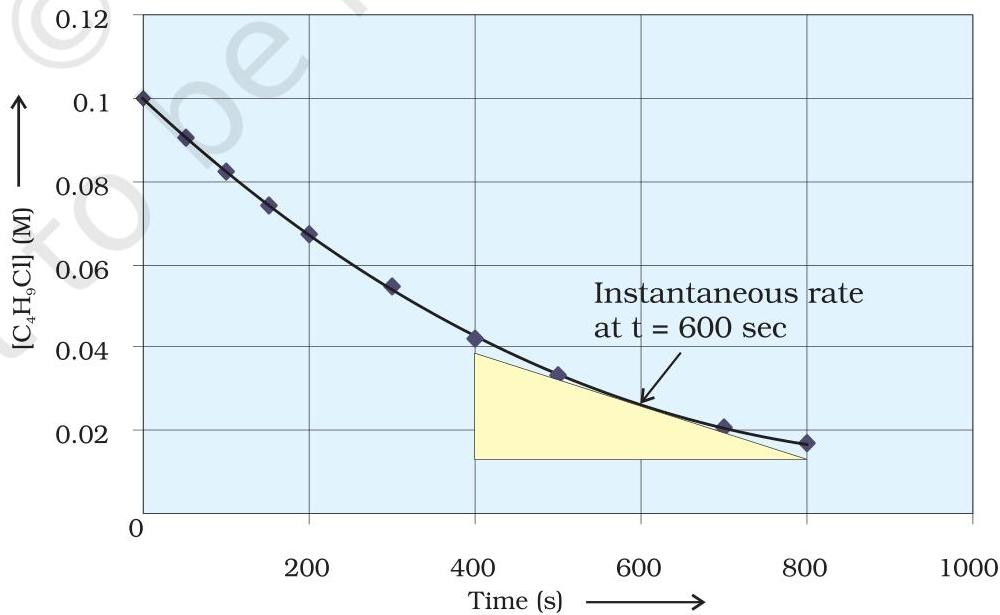
Fig 3.2 Instantaneous rate of hydrolysis of butyl chloride $(\mathrm{C_4H_9Cl})$
The instantaneous rate can be ascertained graphically. This involves constructing a tangent to either the reactant (R) or product (P) concentration-time curve at a specific time point, $t$, and subsequently computing its gradient (
refer to Fig. 3.1). For instance, within the context of problem 3.1, the instantaneous rate ($r_{\mathrm{inst}}$) at 600 seconds can be derived by charting the concentration of butyl chloride against time. A tangent is then precisely drawn, touching the curve at the 600 s mark (as illustrated in Fig. 3.2).
The gradient of this tangent directly corresponds to the instantaneous reaction rate.
$\text{So, } r_{\text{inst}} \text{ at } 600 \text{ s} = - \left( \frac{0.0165 - 0.037}{(800 - 400) \text{ s}} \right) \text{ mol } \mathrm{L}^{-1} = 5.12 \times 10^{-5} \text{ mol } \mathrm{L}^{-1} \text{ s}^{-1}$
$\text{At } t = 250 \text{ s} \quad r_{\text{inst}} = 1.22 \times 10^{-4} \text{ mol } \mathrm{L}^{-1} \text{ s}^{-1}$
$t = 350 \text{ s} \quad r_{\text{inst}} = 1.0 \times 10^{-4} \text{ mol } \mathrm{L}^{-1} \text{ s}^{-1}$
$t = 450 \text{ s} \quad r_{\text{inst}} = 6.4 \times 10^{-5} \text{ mol } \mathrm{L}^{-1} \text{ s}^{-1}$
Let us now examine a chemical reaction:
$\mathrm{Hg}(l) + \mathrm{Cl}_2(g) \rightarrow \mathrm{HgCl}_2(s)$
In cases where the stoichiometric coefficients for both reactants and products are identical, the reaction rate is expressed as follows:
$\text{Rate of reaction} = - \frac{\Delta [\mathrm{Hg}]}{\Delta t} = - \frac{\Delta [\mathrm{Cl}_2]}{\Delta t} = \frac{\Delta [\mathrm{HgCl}_2]}{\Delta t}$
This implies that the rate at which any reactant is consumed is equivalent to the rate at which any product is formed. However, in the subsequent reaction, two moles of hydrogen iodide (HI) undergo decomposition, yielding one mole of hydrogen ($\mathrm{H}_2$) and one mole of iodine ($\mathrm{I}_2$).
$2 \mathrm{HI}(g) \rightarrow \mathrm{H}_2(g) + \mathrm{I}_2(g)$
To articulate the rate of reactions in which the stoichiometric coefficients of reactants or products deviate from unity, the rate of consumption of any reactant or the rate of formation of any product must be normalized by dividing it by its corresponding stoichiometric coefficient. Given that the rate at which HI is consumed is twice the rate at which either $\mathrm{H}_2$ or $\mathrm{I}_2$ is generated, the term $\Delta[\mathrm{HI}]$ is divided by 2 to establish equivalence. The rate of this specific reaction is therefore defined as:
$\text{Rate of reaction} = - \frac{1}{2} \frac{\Delta [\mathrm{HI}]}{\Delta t} = \frac{\Delta [\mathrm{H}_2]}{\Delta t} = \frac{\Delta [\mathrm{I}_2]}{\Delta t}$
Analogously, for the reaction:
$5 \mathrm{Br}^-(\mathrm{aq}) + \mathrm{BrO}_3^-(\mathrm{aq}) + 6 \mathrm{H}^+(\mathrm{aq}) \rightarrow 3 \mathrm{Br}_2(\mathrm{aq}) + 3 \mathrm{H}_2\mathrm{O}(l)$
$\text{Rate} = -\tfrac{1}{5},\tfrac{\Delta[\mathrm{Br}^-]}{\Delta t} = -\tfrac{\Delta[\mathrm{BrO}_3^-]}{\Delta t} = -\tfrac{1}{6},\tfrac{\Delta[\mathrm{H}^+]}{\Delta t} = \tfrac{1}{3},\tfrac{\Delta[\mathrm{Br}_2]}{\Delta t} = \tfrac{1}{3},\tfrac{\Delta[\mathrm{H}_2\mathrm{O}]}{\Delta t}$
In the context of a gaseous reaction occurring at a constant temperature, the concentration of a given species bears a direct proportionality to its partial pressure. Consequently, the reaction rate can alternatively be formulated as the rate of alteration in the partial pressure of either a reactant or a product.
Chemical Kinetics
Example 3.2
An investigation into the decomposition of dinitrogen pentoxide ($\mathrm{N}_2\mathrm{O}_5$) in a carbon tetrachloride ($\mathrm{CCl}_4$) solvent at a temperature of 318 K was conducted by observing the changes in $\mathrm{N}_2\mathrm{O}_5$ concentration within the solution. The initial concentration of $\mathrm{N}_2\mathrm{O}_5$ was recorded as $2.33,\mathrm{mol},\mathrm{L}^{-1}$, subsequently decreasing to $2.08,\mathrm{mol},\mathrm{L}^{-1}$ over a period of 184 minutes. This reaction proceeds according to the following stoichiometric representation:
$2,\mathrm{N}_2\mathrm{O}_5,(\mathrm{g}) \rightarrow 4,\mathrm{NO}_2,(\mathrm{g}) + \mathrm{O}_2,(\mathrm{g})$
Determine the average reaction rate, expressing it in units of hours, minutes, and seconds. Additionally, ascertain the rate at which nitrogen dioxide ($\mathrm{NO}_2$) is generated throughout this interval.
Solution
The average rate of reaction is computed as:
$\text{Average Rate} = \frac{1}{2}\left{-\frac{\Delta[\mathrm{N}_2\mathrm{O}_5]}{\Delta t}\right} = -\frac{1}{2}\left[\frac{(2.08 - 2.33),\mathrm{mol},\mathrm{L}^{-1}}{184,\mathrm{min}}\right]$
$\begin{array}{l} = 6.79 \times 10^{-4},\mathrm{mol},\mathrm{L}^{-1}/\mathrm{min} = (6.79 \times 10^{-4},\mathrm{mol},\mathrm{L}^{-1},\mathrm{min}^{-1}) \times (60,\mathrm{min}/1,\mathrm{h}) \ = 4.07 \times 10^{-2},\mathrm{mol},\mathrm{L}^{-1}/\mathrm{h} \ = 6.79 \times 10^{-4},\mathrm{mol},\mathrm{L}^{-1} \times 1,\mathrm{min}/60,\mathrm{s} \ = 1.13 \times 10^{-5},\mathrm{mol},\mathrm{L}^{-1},\mathrm{s}^{-1} \ \end{array}$
Recall that the reaction rate can also be expressed as:
$\text{Rate} = \frac{1}{4} \left{\frac{\Delta[\mathrm{NO}_2]}{\Delta t}\right}$
$\frac{\Delta[\mathrm{NO}_2]}{\Delta t} = 6.79 \times 10^{-4} \times 4,\mathrm{mol},\mathrm{L}^{-1},\mathrm{min}^{-1} = 2.72 \times 10^{-3},\mathrm{mol},\mathrm{L}^{-1},\mathrm{min}^{-1}$
Intext Questions
3.1 For the reaction $\text{R} \rightarrow \text{P}$, the concentration of a reactant changes from $0.03\text{ M}$ to $0.02\text{ M}$ in $25\text{ minutes}$. Calculate the average rate of reaction using units of time both in minutes and seconds.
3.2 In a reaction,
$2\text{A} \rightarrow \text{Products}$
the concentration of A decreases from $0.5\text{ mol L}^{-1}$ to $0.4\text{ mol L}^{-1}$ in $10\text{ minutes}$. Calculate the rate during this interval.
3.2 Factors Influencing Rate of a Reaction
The speed at which a chemical reaction proceeds is contingent upon various experimental parameters, including the concentration of the reacting species (or pressure for gaseous reactants), the ambient temperature, and the presence of a catalyst.
3.2.1 Dependence of Rate on Concentration
At a specific temperature, the pace of a chemical transformation can be influenced by the concentrations of one or more participating reactants and products. The mathematical description of a reaction's rate, expressed with respect to the concentrations of its reactants, is defined as the rate law. This concept is also frequently referred to as the rate equation or rate expression.
3.2.2 Rate Expression and Rate Constant
The data presented in Table 3.1 unequivocally demonstrates that a reaction's velocity diminishes over time, correlating with the reduction in reactant concentrations. Conversely, an increase in reactant concentrations typically leads to an acceleration of reaction rates. Consequently, the reaction rate is fundamentally contingent upon the concentrations of the reactants.
Consider a general reaction
$\mathrm {a A} + \mathrm {b B} \rightarrow \mathrm {c C} + \mathrm {d D}$
where a, b, c, and d represent the respective stoichiometric coefficients for the reactants and products involved.
The rate expression for this reaction is
$\text {R a t e} \propto [ \mathrm {A} ] ^ {\mathrm {x}} [ \mathrm {B} ] ^ {\mathrm {y}} \tag {3.4}$
where the exponents $x$ and $y$ are not necessarily equivalent to the stoichiometric coefficients (a and b) of the reactants. This equation can alternatively be expressed as
$\text {R a t e} = k [ \mathrm {A} ] ^ {\mathrm {x}} [ \mathrm {B} ] ^ {\mathrm {y}} \tag {3.4a}$
$- \frac {\mathrm {d} [ \mathrm {R} ]}{\mathrm {d} t} = k [ \mathrm {A} ] ^ {\mathrm {x}} [ \mathrm {B} ] ^ {\mathrm {y}} \tag {3.4b}$
The mathematical representation found in equation (3.4b) is designated as the differential rate equation, wherein $k$ signifies a proportionality constant known as the rate constant. An equation such as (3.4), which establishes a relationship between a reaction's speed and the concentrations of its reactants, is termed a rate law or rate expression. Hence, a rate law constitutes an expression where the reaction rate is articulated using the molar concentrations of reactants, with each concentration term elevated to an exponent that may or may not correspond to the stoichiometric coefficient of the respective reacting species in a balanced chemical equation. For instance:
$2 \mathrm {N O} (\mathrm {g}) + \mathrm {O} _ {2} (\mathrm {g}) \rightarrow 2 \mathrm {N O} _ {2} (\mathrm {g})$
The reaction rate can be empirically determined as a function of initial concentrations. This is typically achieved either by maintaining the concentration of one reactant at a constant level while systematically varying the concentration of the other, or by concurrently altering the concentrations of both reactants. The subsequent findings are tabulated (Table 3.2).
Table 3.2: Initial rate of formation of $\mathrm{NO}_2$
| Experiment | Initial [NO]/ mol L⁻¹ | Initial [O₂]/ mol L⁻¹ | Initial rate of formation of NO₂/ mol L⁻¹s⁻¹ |
|---|---|---|---|
| 1. | 0.30 | 0.30 | 0.096 |
| 2. | 0.60 | 0.30 | 0.384 |
| 3. | 0.30 | 0.60 | 0.192 |
| 4. | 0.60 | 0.60 | 0.768 |
Upon examining the experimental data presented, it becomes apparent that a twofold increase in the initial concentration of NO, while maintaining a constant $\mathrm{O}_2$ concentration, leads to a quadrupling of the initial reaction rate, specifically from 0.096 to $0.384,\mathrm{mol},\mathrm{L}^{-1},\mathrm{s}^{-1}$. This observation signifies that the reaction rate is proportional to the square of the NO concentration. Conversely, when the NO concentration is held constant and the $\mathrm{O}_2$ concentration is doubled, the reaction rate also doubles, thereby indicating a first-
order dependence on the $\mathrm{O}_2$ concentration. Consequently, the rate equation for this particular reaction can be expressed as:
$\text {R a t e} = k [ \mathrm {N O} ] ^ {2} [ \mathrm {O} _ {2} ]$
The differential form of this rate expression is given as
$- \frac {\mathrm {d} [ \mathrm {R} ]}{\mathrm {d} t} = k [ \mathrm {N O} ] ^ {2} \left[ \mathrm {O} _ {2} \right]$
It is noteworthy that, for the specific reaction under consideration, the exponents appearing in the experimentally derived rate equation correspond precisely to the stoichiometric coefficients of the respective reactants in the balanced chemical equation.
Some other examples are given below:
| Reaction | Experimental rate expression |
|---|---|
| 1. CHCl${3}$ + Cl${2}$ → CCl$_{4}$ + HCl | Rate = k [CHCl${3}$] [Cl${2}$]$^{1/2}$ |
| 2. CH${3}$COOC${2}$H${5}$ + H${2}$O → CH${3}$COOH + C${2}$H$_{5}$OH | Rate = k [CH${3}$COOC${2}$H${5}$] [H${2}$O]$^{0}$ |
Conversely, in these subsequent reactions, the exponents associated with the concentration terms do not align with their corresponding stoichiometric coefficients. This leads to the conclusion that:
The rate law governing any chemical reaction is not amenable to theoretical prediction solely from an inspection of the balanced chemical equation; rather, its determination necessitates empirical investigation.
3.2.3 Order of a Reaction
Within the rate equation (3.4), presented as
$\text{Rate} = k [\mathrm{A}]^{\mathrm{x}} [\mathrm{B}]^{\mathrm{y}}$
the exponents x and y quantify the dependency of the reaction rate on the concentrations of reactants A and B, respectively. The sum of these exponents, denoted as x + y in expression (3.4), defines the overall order of the reaction, while x and y separately indicate the order relative to reactant A and reactant B.
Consequently, the order of a chemical reaction is defined as the aggregate of the powers to which the reactant concentrations are raised within the rate law equation.
The order of a reaction may assume integer values such as 0, 1, 2, or 3, and can also be fractional. A reaction characterized as zero-order exhibits a rate that is unaffected by alterations in reactant concentrations.
Example 3.3
Determine the total order for a reaction described by the following rate expressions:
(a) Rate = k [A]$^{1/2}$ [B]$^{3/2}$ (b) Rate = k [A]$^{3/2}$ [B]$^{-1}$ (a) order = x + y So order = 1/2 + 3/2 = 2, i.e., second order (b) order = 3/2 + (-1) = 1/2, i.e., half order.
A stoichiometric chemical equation often fails to accurately depict the actual pathway of a reaction, as most reactions do not proceed in a single step. Reactions that occur via a solitary step are termed elementary reactions. Conversely, when products are formed through a series of elementary reactions, collectively known as a mechanism, these are designated as complex reactions.
Such reactions encompass consecutive reactions (for instance, the oxidation of ethane to $\mathrm{CO}{2}$ and $\mathrm{H}{2}\mathrm{O}$ involves several intermediate stages where alcohol, aldehyde, and acid are generated), reverse reactions, and side reactions (such as the nitration of phenol, which produces both $o$-nitrophenol and $p$-nitrophenol).
Units of rate constant
Considering a generic chemical reaction:
$\mathrm{aA} + \mathrm{bB} \rightarrow \mathrm{cC} + \mathrm{dD}$
The rate expression is typically given as:
$\mathrm{Rate} = k , [\mathrm{A}]^{\mathrm{x}} , [\mathrm{B}]^{\mathrm{y}}$
Here, the overall reaction order, denoted by $n$, is the sum of the exponents $x$ and $y$ ($n = x + y$).
The rate constant $k$ can thus be expressed as:
$\begin{array}{l} k = \frac{\mathrm{Rate}}{[\mathrm{A}]^{\mathrm{x}} , [\mathrm{B}]^{\mathrm{y}}} \ = \frac{\text{concentration}}{\text{time}} \times \frac{1}{(\text{concentration})^{\mathrm{n}}} \quad (\text{where } [\mathrm{A}] = [\mathrm{B}]) \end{array}$
Employing the SI units of mol L$^{-1}$ for concentration and seconds (s) for time, the corresponding units for the rate constant $k$ across various reaction orders are delineated in Table 3.3.
Table 3.3: Units of rate constant
| Reaction | Order | Units of rate constant |
|---|---|---|
| Zero order reaction | 0 | $\frac{\mathrm{mol} , \mathrm{L}^{-1}}{\mathrm{s}} \times \frac{1}{(\mathrm{mol} , \mathrm{L}^{-1})^{\mathrm{0}}} = \mathrm{mol} , \mathrm{L}^{-1} , \mathrm{s}^{-1}$ |
| First order reaction | 1 | $\frac{\mathrm{mol} , \mathrm{L}^{-1}}{\mathrm{s}} \times \frac{1}{(\mathrm{mol} , \mathrm{L}^{-1})^{\mathrm{1}}} = \mathrm{s}^{-1}$ |
| Second order reaction | 2 | $\frac{\mathrm{mol} , \mathrm{L}^{-1}}{\mathrm{s}} \times \frac{1}{(\mathrm{mol} , \mathrm{L}^{-1})^{\mathrm{2}}} = \mathrm{mol}^{-1} , \mathrm{L} , \mathrm{s}^{-1}$ |
Determine the reaction order corresponding to each of the subsequent rate constant values. Example 3.4
(i) $k = 2.3 \times 10^{-5} , \mathrm{L} , \mathrm{mol}^{-1} , \mathrm{s}^{-1}$ (ii) $k = 3 \times 10^{-4} , \mathrm{s}^{-1}$ (i) Given that the units for a second-order rate constant are $\mathrm{L} , \mathrm{mol}^{-1} , \mathrm{s}^{-1}$, the provided rate constant $k = 2.3 \times 10^{-5} , \mathrm{L} , \mathrm{mol}^{-1} , \mathrm{s}^{-1}$ signifies a reaction of the second order. (ii) Since $\mathrm{s}^{-1}$ is the characteristic unit for a first-order rate constant, a value of $k = 3 \times 10^{-4} , \mathrm{s}^{-1}$ indicates a first-order reaction.
3.2.4 Molecularity of a Reaction
Molecularity, a distinct characteristic of a reaction, contributes to deciphering its mechanistic pathway. It is defined as the count of reactant entities (be they atoms, ions, or molecules) participating in an elementary reaction that are required to undergo simultaneous collision for the chemical transformation to occur. A reaction is classified as unimolecular if it involves a single reacting species, such as in the decomposition of ammonium nitrite.
$\text{NH}{4}\text{NO}{2} \rightarrow \text{N}{2} + 2\text{H}{2}\text{O}$
In contrast, bimolecular reactions necessitate the concurrent collision of two distinct species; an illustrative instance is the dissociation of hydrogen iodide.
$2\text{HI} \rightarrow \text{H}{2} + \text{I}{2}$
Reactions designated as trimolecular, or alternatively termolecular, are characterized by the simultaneous interaction of three reactant species, for example,
$2\text{NO} + \text{O}{2} \rightarrow 2\text{NO}{2}$
The likelihood of more than three molecules concurrently colliding and reacting is exceedingly low. Consequently, reactions possessing a molecularity of three are exceptionally uncommon and proceed at a sluggish pace.
Thus, it becomes apparent that intricate reactions whose stoichiometric representation includes over three molecules invariably unfold through a sequence of multiple steps.
$\text{KClO}{3} + 6\text{FeSO}{4} + 3\text{H}{2}\text{SO}{4} \rightarrow \text{KCl} + 3\text{Fe}{2}(\text{SO}{4}){3} + 3\text{H}{2}\text{O}$
Although this reaction superficially appears to be tenth order, it is, in fact, a second-order reaction. This observation indicates that the reaction proceeds via multiple stages. The query then arises: which particular stage governs the rate of the entire reaction? This question can be addressed by examining the reaction mechanism. Analogously, a relay race team's prospects of victory are contingent upon its slowest member. In a similar vein, the overall reaction rate is dictated by the slowest elementary step within the reaction sequence, which is termed the rate-determining step. Let us consider the decomposition of hydrogen peroxide, which is catalyzed by iodide ions in an alkaline environment.
$2\text{H}{2}\text{O}{2} - \frac{\Gamma}{\text{Alkaline medium}} \rightarrow 2\text{H}{2}\text{O} + \text{O}{2}$
The rate equation for this reaction is found to be
$\text{Rate} = \frac{-\text{d}\left[\text{H}{2}\text{O}{2}\right]}{\text{d}t} = k\left[\text{H}{2}\text{O}{2}\right]\left[\Gamma\right]$
This reaction exhibits first-order kinetics concerning both $\text{H}{2}\text{O}{2}$ and $\Gamma$. Empirical data indicate that this reaction proceeds via a two-step mechanism:
(1) $\text{H}{2}\text{O}{2} + \Gamma \rightarrow \text{H}{2}\text{O} + \text{IO}^{-}$ (2) $\text{H}{2}\text{O}{2} + \text{IO}^{-} \rightarrow \text{H}{2}\text{O} + \Gamma + \text{O}_{2}$
Each of these steps constitutes a bimolecular elementary reaction. The species $\text{IO}^{-}$ is designated as an intermediate, given its generation during the reaction process but its absence from the final overall balanced chemical equation. The initial step, characterize
d by its slower pace, functions as the rate-determining step. Consequently, the rate at which this intermediate forms dictates the overall rate of this reaction.
From the preceding discourse, we can summarize the following key points:
- The order of a reaction is an experimentally determined value that can be zero or even a fractional number, whereas molecularity cannot be zero or a non-integer. 2. Reaction order is applicable to both elementary and complex reactions, while molecularity is exclusively defined for elementary reactions. For complex reactions, the concept of molecularity is not pertinent.
(iii) For complex reactions, the order is dictated by the slowest step, and the molecularity of this rate-determining step is equivalent to the overall reaction order.
Intext Questions
3.3 Consider a reaction, $A + B \rightarrow$ Product; for which the rate law is expressed as, $r = k[A]^{1/2}[B]^2$. Determine the overall order of this reaction.
3.4 The transformation of molecules $X$ into $Y$ adheres to second-order kinetics. If the concentration of $X$ is augmented threefold, how will this influence the rate at which $Y$ is produced?
3.3 Integrated Rate Equations
As previously discussed, the relationship between reaction rate and reactant concentrations is termed the differential rate equation. Determining the instantaneous rate is often impractical, as it involves calculating the slope of the tangent at a specific time point $t$ on a concentration-versus-time plot (Fig. 3.1). This complexity hinders the straightforward determination of the rate law and, by extension, the reaction order. To overcome this challenge, we can integrate the differential rate equation, thereby establishing a relationship between directly observable experimental data—specifically, concentrations measured at various times—and the rate constant.
The integrated rate equations are distinct for reactions exhibiting different orders. Our examination will be limited to deriving these equations for zero-order and first-order chemical reactions exclusively.
3.3.1 Zero Order Reactions
A reaction is classified as zero order when its rate is independent of the reactant concentration, specifically, proportional to the zeroth power of the reactant's concentration. Let's consider a generic chemical transformation:
$R \rightarrow P$
The differential rate law for this process is given by:
$\text{Rate} = -\frac{d[R]}{dt} = k[R]^0$
Given that any non-zero quantity raised to the power of zero equals one, the expression simplifies to:
$\text{Rate} = -\frac{d[R]}{dt} = k \times 1$
This further implies:
$d[R] = -k dt$
Upon integration of both sides, we obtain:
$[R] = -kt + I \tag{3.5}$
Here, $I$ represents the integration constant.
To determine the value of $I$, we apply the initial condition: at time $t = 0$, the reactant concentration $[R]$ is equal to its initial concentration, denoted as $[R]_0$.
Substituting these conditions into equation (3.5):
$[R]_0 = -k \times 0 + I$
Which simplifies to:
$[R]_0 = I$
By substituting this determined value of $I$ back into equation (3.5), we derive the integrated rate law for a zero-order reaction:
$[R] = -kt + [R]_0 \tag{3.6}$
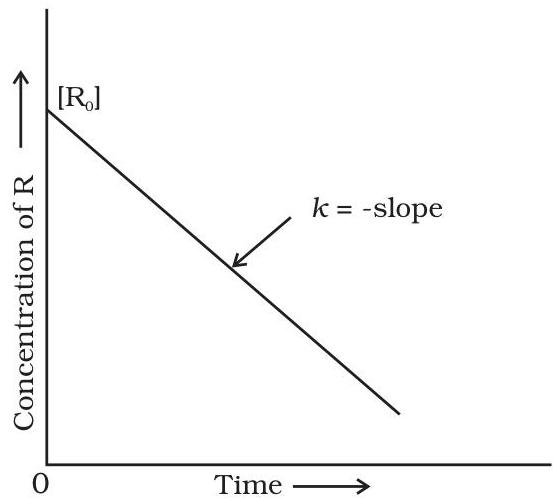
Fig. 3.3: Variation in the concentration vs time plot for a zero order reaction
When equation (3.6) is juxtaposed with the standard linear equation, $y = mx + c$, it becomes evident that a plot of $[R]$ versus $t$ will yield a linear relationship (as illustrated in Fig. 3.3). This straight line will possess a slope equivalent to $-k$ and an intercept corresponding to $[R]_0$.
Rearranging equation (3.6) allows us to isolate the rate constant, $k$:
$k = \frac{[R]_0 - [R]}{t} \tag{3.7}$
While zero-order reactions are not frequently encountered, they manifest under specific circumstances. Notable instances include certain enzyme-catalyzed processes and reactions occurring on metallic surfaces. A prime illustration is the decomposition of gaseous ammonia on a heated platinum surface, which exhibits zero-order kinetics at elevated pressures.
$2NH_3(g) \xrightarrow[Pt\ catalyst]{1130K} N_2(g) + 3H_2(g)$
The rate expression for this reaction is:
$\text{Rate} = k \left[NH_3\right]^0 = k$
In this particular reaction, the platinum metal functions as a catalytic agent. Under high-pressure conditions, the surface of the metal becomes entirely covered by reactant gas molecules. Consequently, subsequent alterations in the reaction environment do not change the quantity of ammonia adsorbed on the catalyst surface, thereby rendering the reaction rate independent of the ammonia concentration. Another example of a zero-order reaction is the thermal decomposition of hydrogen iodide on a gold surface.
3.3.2 First Order Reactions
Reactions categorized as first-order exhibit a reaction rate directly proportional to the reactant R's concentration raised to the first power. Consider, for instance, a general reaction:
$R \rightarrow P$
$\text{Rate} = -\frac{d[R]}{dt} = k[R]$
$\text{or} \quad \frac{d[R]}{[R]} = -k dt$
Integrating this equation yields:
$\ln [R] = -kt + I \tag{3.8}$
The variable $I$ represents the integration constant, which can be readily determined.
At the reaction's initiation ($t = 0$), the reactant concentration $R$ is denoted as $[R]_0$, signifying its initial value.
Consequently, substituting these initial conditions into equation (3.8) yields:
$\ln [R]_0 = -k \times 0 + I$
$\ln [R]_0 = I$
By substituting this derived value of $I$ back into equation (3.8):
$\ln[R] = -kt + \ln[R]_0 \tag{3.9}$
Rearranging the terms of this equation leads to:
$\ln \frac {[ \mathrm {R} ]}{[ \mathrm {R} ] _ {0}} = - k t$
$\text {or} \ k = \frac {1}{t} \ln \frac {\left[ R \right] _ {0}}{[ R ]} \tag {3.10}$
Considering equation (3.8) at a specific time $t_1$:
$\ln [ R ] _ {1} = - k t _ {1} + \ln [ R ] _ {0} \tag {3.11}$
Similarly, at a subsequent time $t_2$:
$\ln \left[ R \right] _ {2} = - k t _ {2} + \ln \left[ R \right] _ {0} \tag {3.12}$
Here, $[R]_1$ and $[R]_2$ denote the reactant concentrations observed at times $t_1$ and $t_2$, respectively.
Subtracting equation (3.12) from equation (3.11) provides:
$\ln \left[ R \right] _ {1} - \ln \left[ R \right] _ {2} = - k t _ {1} - (- k t _ {2})$
$\ln \frac {[ \mathrm {R} ] _ {1}}{[ \mathrm {R} ] _ {2}} = k (t _ {2} - t _ {1})$
$k = \frac {1}{\left(t _ {2} - t _ {1}\right)} \ln \frac {[ \mathrm {R} ] _ {1}}{[ \mathrm {R} ] _ {2}} \tag {3.13}$
Equation (3.9) can alternatively be expressed as:
$\ln \frac {[ \mathrm {R} ]}{[ \mathrm {R} ] _ {0}} = - k t$
Applying the antilogarithm to both sides of the equation:
$[ \mathrm {R} ] = [ \mathrm {R} ] _ {0} \mathrm {e} ^ {- k t} \tag {3.14}$
By drawing an analogy between equation (3.9) and the linear equation $y = mx + c$, a plot of $\ln [R]$ versus $t$ (as depicted in Fig. 3.4) will yield a linear relationship. This line will possess a slope equal to $-k$ and an intercept corresponding to $\ln [R]_0$.
The integrated rate equation for a first-order reaction (equation 3.10) can also be presented in the following form:
$k = \frac {2.303}{t} \log \frac {[ R ] _ {0}}{[ R ]} \tag {3.15}$
$\log \frac {[ R ] _ {0}}{[ R ]} = \frac {k t}{2.303}$
When a graph is constructed plotting $\log [R]_0 / [R]$ against time $t$ (refer to Fig. 3.5),
the slope $= k / 2.303$
The hydrogenation of ethene serves as a representative instance of a first-order reaction.
$\mathrm {C} _ {2} \mathrm {H} _ {4} (\mathrm {g}) + \mathrm {H} _ {2} (\mathrm {g}) \rightarrow \mathrm {C} _ {2} \mathrm {H} _ {6} (\mathrm {g})$
$\text {Rate} = k \left[ \mathrm {C} _ {2} \mathrm {H} _ {4} \right]$
Both naturally occurring and artificially induced radioactive decay processes of unstable atomic nuclei invariably conform to first-order kinetics.
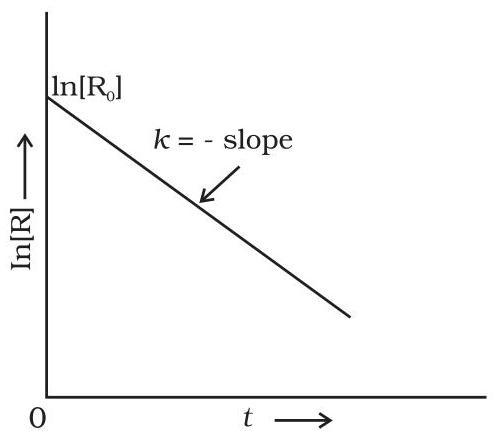
Fig. 3.4: A plot between $\ln [R]$ and $t$ for a first order reaction
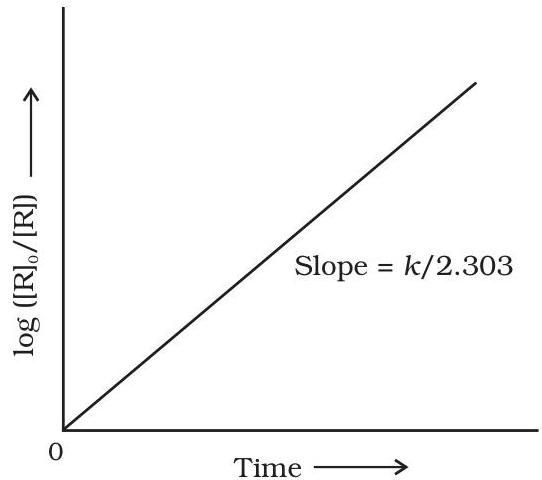
Fig. 3.5: Plot of $\log [R]_0 / [R]$ vs time for a first order reaction
${ } _ { 8 8 } ^ { 2 2 6 } \mathrm {R a} \rightarrow { } _ { 2 } ^ { 4 } \mathrm {H e} + { } _ { 8 6 } ^ { 2 2 2 } \mathrm {R n}$
$\mathrm {Rate} = k [ \mathrm {Ra} ]$
The decomposition processes of $\mathrm{N}_2\mathrm{O}_5$ and $\mathrm{N}_2\mathrm{O}$ serve as additional illustrations of reactions exhibiting first-order kinetics.
🧪 Example 3.5
For the first-order reaction
$\mathrm{N}_2\mathrm{O}_5(\mathrm{g}) \rightarrow 2\mathrm{NO}_2(\mathrm{g}) + \frac{1}{2}\mathrm{O}_2(\mathrm{g})$
the initial concentration of $\mathrm{N}_2\mathrm{O}_5$ at $318\mathrm{K}$ was recorded as $1.24 \times 10^{-2}\mathrm{mol}\mathrm{L}^{-1}$. Subsequently, after an interval of 60 minutes, its concentration diminished to $0.20 \times 10^{-2}\mathrm{mol}\mathrm{L}^{-1}$. Determine the reaction's rate constant at $318\mathrm{K}$ .
Solution: For a first order reaction
$\begin{array}{l} \log \frac {[ R ] _ {1}}{[ R ] _ {2}} = \frac {k (t _ {2} - t _ {1})}{2 . 3 0 3} \ k = \frac {2 . 3 0 3}{\left(t _ {2} - t _ {1}\right)} \log \frac {1 . 2 4 \times 1 0 ^ {- 2} \mathrm {mol L} ^ {- 1}}{0 . 2 0 \times 1 0 ^ {- 2} \mathrm {mol L} ^ {- 1}} \ = \frac {2 . 3 0 3}{6 0} \log 6. 2 \min ^ {- 1} \ k = 0. 0 3 0 4 \mathrm {min} ^ {- 1} \ \end{array}$
Consider the following representative first-order gas-phase reaction:
$\mathrm {A} (\mathrm {g}) \rightarrow \mathrm {B} (\mathrm {g}) + \mathrm {C} (\mathrm {g})$
Let $p_{\mathrm{i}}$ denote the initial partial pressure of reactant A, and $p_{\mathrm{t}}$ represent the cumulative total pressure observed at a given time 't'. The integrated rate equation pertinent to such a reaction can be formulated as follows:
The total pressure, $p_{\mathrm{t}}$, is defined as the sum of the partial pressures of the constituent gases: $p_{\mathrm{t}} = p_{\mathrm{A}} + p_{\mathrm{B}} + p_{\mathrm{C}}$ (where pressures are in consistent units).
$p_{\mathrm{A}}$, $p_{\mathrm{B}}$, and $p_{\mathrm{C}}$ denote the partial pressures corresponding to species A, B, and C, respectively.
Assuming $\mathbf{x}$ atm signifies the reduction in the partial pressure of A at time $t$, and given that one mole each of B and C is generated, the concomitant increase in the partial pressures of B and C will likewise be $\mathbf{x}$ atm for each.
$\mathrm{A}(\mathrm{g}) \quad \rightarrow \quad \mathrm{B}(\mathrm{g}) \quad + \quad \mathrm{C}(\mathrm{g})$
At $t = 0$: $p_{\mathrm{i}}$ atm 0 atm 0 atm At time $t$: $(p_{\mathrm{i}} - \mathbf{x})$ atm $\mathbf{x}$ atm $\mathbf{x}$ atm
where $p_{\mathrm{i}}$ represents the initial pressure at the reaction's onset, specifically at $t=0$.
$\begin{array}{l} p_{\mathrm{t}} = (p_{\mathrm{i}} - \mathbf{x}) + \mathbf{x} + \mathbf{x} = p_{\mathrm{i}} + \mathbf{x} \ \mathbf{x} = p_{\mathrm{t}} - p_{\mathrm{i}} \end{array}$
Consequently, the partial pressure of A at time $t$, denoted $p_{\mathrm{A}}$, can be expressed as: $p_{\mathrm{A}} = p_{\mathrm{i}} - \mathbf{x}$. Substituting the expression for $\mathbf{x}$, we get $p_{\mathrm{A}} = p_{\mathrm{i}} - (p_{\mathrm{t}} - p_{\mathrm{i}})$.
$= 2 p_{\mathrm{i}} - p_{\mathrm{t}}$
$\begin{array}{l} k = \left(\frac{2.303}{t}\right) \left(\log \frac{p_{\mathrm{i}}}{p_{\mathrm{A}}}\right) \tag{3.16} \ = \frac{2.303}{t} \log \frac{p_{\mathrm{i}}}{(2 p_{\mathrm{i}} - p_{\mathrm{t}})} \end{array}$
The subsequent experimental data were acquired from the first-order thermal decomposition of gaseous $\mathrm{N}_2\mathrm{O}_5$ conducted under conditions of constant volume:
$2 \mathrm{N}_2 \mathrm{O}_5(\mathrm{g}) \rightarrow 2 \mathrm{N}_2 \mathrm{O}_4(\mathrm{g}) + \mathrm{O}_2(\mathrm{g})$
Determine the rate constant for this reaction.
Considering the stoichiometry of the reaction, if the partial pressure of $\mathrm{N}_2\mathrm{O}_5(\mathrm{g})$ diminishes by $2\mathrm{x}$ atmospheres, then the partial pressure of $\mathrm{N}_2\mathrm{O}_4(\mathrm{g})$ will consequently increase by $2\mathrm{x}$ atmospheres, and the partial pressure of $\mathrm{O}_2(\mathrm{g})$ will increase by $\mathrm{x}$ atmospheres. This directly correlates with the molar ratios involved in the decomposition process.
$2 \mathrm{N}_2 \mathrm{O}_5(\mathrm{g}) \quad \rightarrow \quad 2 \mathrm{N}_2 \mathrm{O}_4(\mathrm{g}) \quad + \quad \mathrm{O}_2(\mathrm{g})$
At the initial time, $t = 0$: $\mathrm{N}_2\mathrm{O}_5(\mathrm{g})$: 0.5 atm $\mathrm{N}_2\mathrm{O}_4(\mathrm{g})$: 0 atm $\mathrm{O}_2(\mathrm{g})$: 0 atm
At a subsequent time $t$: $\mathrm{N}_2\mathrm{O}_5(\mathrm{g})$: $(0.5 - 2\mathrm{x})$ atm $\mathrm{N}_2\mathrm{O}_4(\mathrm{g})$: $2\mathrm{x}$ atm $\mathrm{O}_2(\mathrm{g})$: $\mathrm{x}$ atm
$\begin{array}{l} p_{t} = p_{\mathrm{N}_2\mathrm{O}5} + p{\mathrm{N}_2\mathrm{O}4} + p{\mathrm{O}2} \ = (0.5 - 2x) + 2x + x = 0.5 + x \ x = p{t} - 0.5 \end{array}$
$\begin{array}{l} p_{\mathrm{N}_2\mathrm{O}_5} = 0.5 - 2x \
= 0.5 - 2 (p_{\mathrm{t}} - 0.5) = 1.5 - 2 p_{\mathrm{t}} \end{array}$
At $t = 100$ s, the total pressure $p_{\mathrm{t}} = 0.512$ atm.
$p _ {\mathrm {N} _ {2} \mathrm {O} _ {5}} = 1. 5 - 2 \times 0. 5 1 2 = 0. 4 7 6 \mathrm {a t m}$
Using equation (3.16):
$\begin{array}{l} k = \frac {2 . 3 0 3}{t} \log \frac {p _ {\mathrm {i}}}{p _ {\mathrm {A}}} = \frac {2 . 3 0 3}{1 0 0 \mathrm {s}} \log \frac {0 . 5 \mathrm {a t m}}{0 . 4 7 6 \mathrm {a t m}} \ = \frac {2 . 3 0 3}{1 0 0 \mathrm {s}} \times 0. 0 2 1 6 = 4. 9 8 \times 1 0 ^ {- 4} \mathrm {s} ^ {- 1} \ \end{array}$
3.3.3 Half-Life of a Reaction
The half-life of a reaction refers to the duration required for the concentration of a given reactant to decrease to precisely half of its original concentration. This period is denoted by $t_{1/2}$.
For a reaction exhibiting zero-order kinetics, the rate constant is defined by Equation 3.7:
$k = \frac {[ \mathrm {R} ] _ {0} - [ \mathrm {R} ]}{t}$
When the time elapsed is equal to the half-life, $t = t_{1/2}$, the concentration of reactant R is reduced to half its initial value: $[ \mathrm {R} ] = \frac {1}{2} [ \mathrm {R} ] _ {0}$.
Substituting these conditions into the rate constant expression yields:
$k = \frac {[ \mathrm {R} ] _ {0} - 1 / 2 [ \mathrm {R} ] _ {0}}{t _ {1 / 2}}$
Rearranging this equation to solve for the half-life gives:
$t _ {1 / 2} = \frac {[ \mathrm {R} ] _ {0}}{2 k}$
This derivation demonstrates that for a zero-order reaction, the half-life $t_{1/2}$ exhibits a direct proportionality to the initial concentration of the reactants and an inverse proportionality to the rate constant.
For a first-order reaction, the rate constant is expressed as:
$k = \frac {2.303}{t} \log \frac {[ \mathrm {R} ] _ {0}}{[ \mathrm {R} ]} \tag {3.15}$
At the half-life period, $t = t_{1/2}$, the reactant concentration $[ \mathrm {R} ]$ is half of its initial value $[ \mathrm {R} ] _ {0}$:
$[ \mathrm {R} ] = \frac {[ \mathrm {R} ] _ {0}}{2} \tag {3.16}$
Substituting this condition into the integrated rate law results in:
$k = \frac {2.303}{t _ {1 / 2}} \log \frac {[ \mathrm {R} ] _ {0}}{[ \mathrm {R} ] / 2}$
This equation can be rearranged to define the half-life:
$\text {or} \quad t _ {1 / 2} = \frac {2.303}{k} \log 2$
Given that $\log 2 \approx 0.301$, the expression simplifies to:
$t _ {1 / 2} = \frac {2.303}{k} \times 0.301$
Thus, the half-life for a first-order reaction is:
$t _ {1 / 2} = \frac {0.693}{k} \tag {3.17}$
It is observable that for a first-order reaction, the half-life period remains invariant; that is, it does not depend on the initial concentration of the participating chemical species. Consequently, the half-life of a first-order reaction can be directly computed from its rate constant, and conversely.
To summarize, for a zero-order reaction, $t_{1/2} \propto [R]0$, whereas for a first-order reaction, $t{1/2}$ is independent of $[R]_0$.
🧪 Example 3.7
Consider a first-order reaction characterized by a rate constant $k = 5.5 \times 10^{-14} , \text{s}^{-1}$.
Determine the half-life of this reaction.
The half-life for a first-order reaction is calculated using the formula:
$t_{1/2} = \frac{0.693}{k}$
Substituting the given rate constant:
$t_{1/2} = \frac{0.693}{5.5 \times 10^{-14} , \text{s}^{-1}} = 1.26 \times 10^{13} , \text{s}$
Demonstrate that for a first-order reaction, the time needed for $99.9%$ completion is precisely 10 times the half-life ($t_{1/2}$) of that reaction.
Upon $99.9%$ completion of the reaction, the remaining concentration of reactant $[R]$ is given by $[R] = [R]_0 - 0.999[R]_0$.
Applying the integrated rate law for a first-order reaction:
$\begin{array}{l} k = \frac{2.303}{t} \log \frac{[R]_0}{[R]} \ = \frac{2.303}{t} \log \frac{[R]_0}{[R]_0 - 0.999[R]_0} = \frac{2.303}{t} \log 10^3 \ t = 6.909 / k \ \end{array}$
For the half-life of the reaction, we use its known expression:
$\begin{array}{l} t_{1/2} = 0.693 / k \ \frac{t}{t_{1/2}} = \frac{6.909}{k} \times \frac{k}{0.693} = 10 \ \end{array}$
Table 3.4 provides a concise summary of the mathematical characteristics associated with the integrated rate laws for zero and first-order reactions.
Table 3.4: Integrated Rate Laws for the Reactions of Zero and First Order
Chemical Kinetics
The effective order of a reaction can be influenced by specific conditions. Many reactions, while inherently of a higher order, may nonetheless adhere to a first-order rate law. Consider, for instance, the hydrolysis of ethyl acetate, which involves a chemical interaction between ethyl acetate and water. Intrinsically, this process is a second-order reaction, with its rate contingent upon the concentrations of both ethyl acetate and water. However, when water is employed in a substantial stoichiometric excess for the hydrolysis, its concentration remains largely invariant throughout the reaction's progression. Consequently, the reaction rate effectively becomes dependent solely on the concentration of ethyl acetate. For example, during the hydrolysis of 0.01 mol of ethyl acetate with 10 mol of water, the quantities of reactants and products present at the commencement (t = 0) and conclusion (t) of the reaction are presented below.
$\begin{array}{c c c c c c}
\mathrm{CH_3COOC_2H_5} & + \mathrm{H_2O} & \xrightarrow{\mathrm{H^+}} & \mathrm{CH_3COOH} & + & \mathrm{C_2H_5OH} \
t = 0 & 0.01 \mathrm{mol} & 10 \mathrm{mol} & 0 \mathrm{mol} & 0 \mathrm{mol} \
t & 0 \mathrm{mol} & 9.99 \mathrm{mol} & 0.01 \mathrm{mol} & 0.01 \mathrm{mol}
\end{array}$
Due to the negligible alteration in the concentration of water over the reaction's duration, the reaction effectively behaves as a first-order process. Such reactions are designated as pseudo first-order reactions.
The inversion of cane sugar represents another example of a pseudo first-order reaction.
$\begin{array}{c c c c c c} \mathrm{C_{12}H_{22}O_{11}} & + & \mathrm{H_2O} & \xrightarrow{\mathrm{H^+}} & \mathrm{C_6H_{12}O_6} & + & \mathrm{C_6H_{12}O_6} \ \text{Cane sugar} & & & & \text{Glucose} & & \text{Fructose} \end{array}$
$\text{Rate} = k \left[ \mathrm{C_{12}H_{22}O_{11}} \right]$
Intext Questions
3.5 A first order reaction has a rate constant $1.15 \times 10^{-3}\text{ s}^{-1}$. How long will $5\text{ g}$ of this reactant take to reduce to $3\text{ g}$?
3.6 Time required to decompose $\text{SO}_2\text{Cl}_2$ to half of its initial amount is $60\text{ minutes}$. If the decomposition is a first order reaction, calculate the rate constant of the reaction.
3.4 Temperature Dependence of the Rate of a Reaction
An elevation in temperature typically leads to an acceleration in the rates of most chemical reactions. A salient illustration is the decomposition of dinitrogen pentoxide ($\mathrm{N_2O_5}$), where the half-life for the reactant's initial quantity is observed to be $12,\mathrm{min}$ at $50^{\circ}\mathrm{C}$, extending to $5,\mathrm{h}$ at $25^{\circ}\mathrm{C}$, and further to 10 days when the temperature is $0^{\circ}\mathrm{C}$. Similarly, it is a common observation that in a solution containing potassium permanganate ($\mathrm{KMnO_4}$) and oxalic acid ($\mathrm{H_2C_2O_4}$), the decolourisation of potassium permanganate proceeds more rapidly at elevated temperatures compared to lower temperatures.
Empirical evidence indicates that for many chemical reactions, an approximate doubling of the rate constant occurs with a $10^{\circ}$ increase in temperature.
The quantitative relationship between temperature and the rate of a chemical reaction is precisely described by the Arrhenius equation (3.18). While initially conceived by the Dutch chemist J.H. van't Hoff, its comprehensive physical rationale and mechanistic interpretation were later furnished by the Swedish chemist Svante Arrhenius.
$k = \mathrm {A e} ^ {- E a / R T} \tag {3.18}$
In this expression, $A$ denotes the Arrhenius factor, alternatively termed the frequency factor or pre-exponential factor, which represents a characteristic constant unique to a given reaction. $R
$ signifies the universal gas constant, and $E_{\mathrm{a}}$ corresponds to the activation energy, typically expressed in joules per mole ($\mathrm{J mol}^{-1}$).
It can be understood clearly using the following simple reaction
$\mathrm {H} _ {2} (\mathrm {g}) + \mathrm {I} _ {2} (\mathrm {g}) \rightarrow 2 \mathrm {H I} (\mathrm {g})$
Arrhenius's hypothesis posits that this reaction proceeds exclusively upon the collision of a hydrogen molecule with an iodine molecule, leading to the formation of a transient, unstable intermediate species (Fig. 3.6). This intermediate persists for an exceedingly brief duration before dissociating to yield two molecules of hydrogen iodide.
The energetic input necessary for the generation of this intermediate, designated as the activated complex (C), is defined as the activation energy $(E_{\mathrm{a}})$. A visual representation of this energetic profile is depicted in Fig. 3.7, which illustrates potential energy as a function of the reaction coordinate. The reaction coordinate itself serves to map the progression of energy transformations as reactants transition into products.
Upon the decomposition of this complex into products, a certain quantity of energy is liberated. Consequently, the ultimate enthalpy change of the reaction is contingent upon the intrinsic properties of both the initial reactants and the resultant products.
It is important to note that the constituent molecules within a reacting system do not possess uniform kinetic energy. Given the inherent difficulty in forecasting the precise trajectory of individual molecules, Ludwig Boltzmann and James Clark Maxwell applied statistical methods to characterize the collective behavior of macroscopic ensembles of molecules. Their framework suggests that the distribution of kinetic energy can be graphically represented by plotting the proportion of molecules $(N_{\mathrm{E}} / N_{\mathrm{T}})$ exhibiting a specific kinetic energy (E) against the kinetic energy itself (Fig. 3.8). In this context, $N_{\mathrm{E}}$ represents the count of molecules possessing energy $E$, while $N_{\mathrm{T}}$ denotes the aggregate number of molecules.
The apex of the distribution curve indicates the kinetic energy that the greatest proportion of molecules possess. The population of molecules diminishes at kinetic energy values that are either above or below this most probable energy.
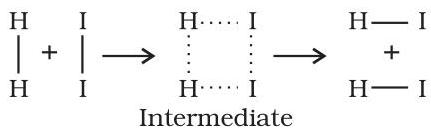
Fig. 3.6: Formation of HI through the intermediate
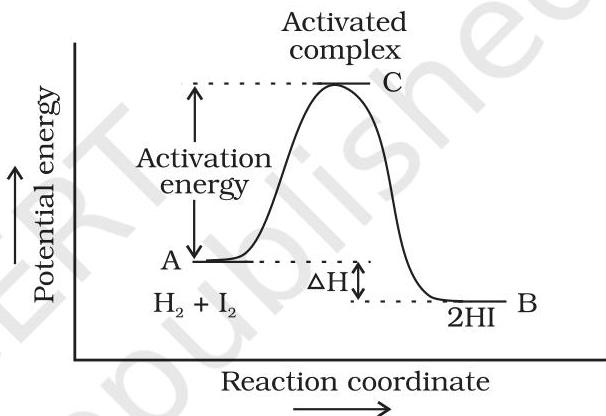
Fig. 3.7: Diagram showing plot of potential energy vs reaction coordinate
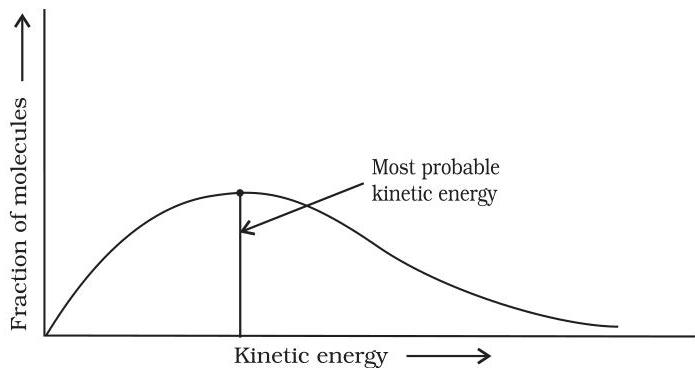
Fig. 3.8: Distribution curve showing energies among gaseous molecules
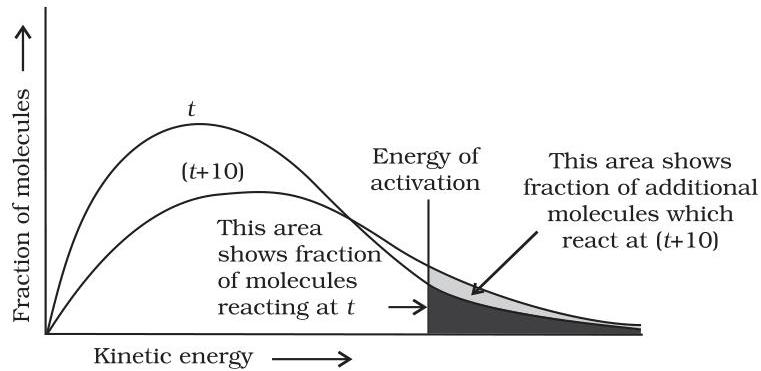
Fig. 3.9: Distribution curve showing temperature dependence of rate of a reaction
Upon an increase in temperature, the curve's maximum point shifts towards greater energy values (Fig. 3.9), and the curve itself expands, or broadens, towards the right. This indicates an augmented proportion of molecules possessing considerably elevated energies. The total area enclosed by the curve must remain invariant, as it signifies a cumulative probability of unity. The activation energy, $E_{\mathrm{a}}$, can be designated on the Maxwell-Boltzmann distribution curve (Fig. 3.9).
Elevating the temperature of a given substance results in an increased proportion of molecules possessing kinetic energies sufficient for collisions exceeding $E_{\mathrm{a}}$. The diagram explicitly illustrates that, for the curve at $(t + 10)$, the region representing the fraction of molecules with energies equal to or surpassing the activation energy is observed to double, consequently leading to a twofold increase in the reaction rate.
Within the Arrhenius equation (3.18), the exponential term, $\mathrm{e}^{-E_{\mathrm{a}} / RT}$, quantifies the proportion of molecules possessing kinetic energy that surpasses $E_{\mathrm{a}}$. Applying the natural logarithm to both members of equation (3.18) yields:
$\ln k = - \frac {E _ {\mathrm {a}}}{R T} + \ln A \tag {3.19}$
A graphical representation of $\ln k$ against $1 / T$ produces a linear relationship, consistent with equation (3.19) and illustrated in Fig. 3.10.
Consequently, analysis of the Arrhenius equation (3.18) reveals that either an elevation in temperature or a reduction in activation energy will lead to an acceleration of the reaction rate and an exponential enhancement of the rate constant.
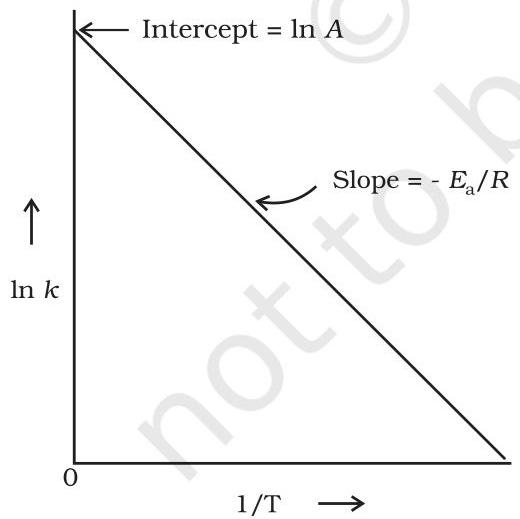
Fig. 3.10: A plot between $\ln k$ and $1 / T$
Referring to Fig. 3.10, the gradient is determined to be $= -\frac{E_{\mathrm{a}}}{R}$ and the y-intercept is $= \ln A$. These derived quantities enable the computation of $E_{\mathrm{a}}$ and $A$. For a temperature $T_{1}$, equation (3.19) is expressed as:
$\ln k _ {1} = - \frac {E _ {\mathrm {a}}}{R T _ {1}} + \ln A \tag {3.20}$
Similarly, at a temperature $T_{2}$, equation (3.19) takes the form:
$\ln k _ {2} = - \frac {E _ {\mathrm {a}}}{R T _ {2}} + \ln A \tag {3.21}$
(given that $A$ remains constant for a particular reaction)
Consider $k_1$ and $k_2$ as the rate constants corresponding to temperatures $T_1$ and $T_2$, respectively.
By performing the subtraction of equation (3.20) from equation (3.21), the following relationship is derived:
$\ln k _ {2} - \ln k _ {1} = \frac {E _ {\mathrm {a}}}{R T _ {1}} - \frac {E _ {\mathrm {a}}}{R T _ {2}}$
$\ln \frac {k _ {2}}{k _ {1}} = \frac {E _ {\mathrm {a}}}{R} \left[ \frac {1}{T _ {1}} - \frac {1}{T _ {2}} \right]$
$\log \frac {k _ {2}}{k _ {1}} = \frac {E _ {\mathrm {a}}}{2 . 3 0 3 R} \left[ \frac {1}{T _ {1}} - \frac {1}{T _ {2}} \right] \tag {3.22}$
$\log \frac {k _ {2}}{k _ {1}} = \frac {E _ {\mathrm {a}}}{2 . 3 0 3 \mathrm {R}} \left[ \frac {T _ {2} - T _ {1}}{T _ {1} T _ {2}} \right]$
🧪 Example 3.8
For a given reaction, the rate constants observed at 500 K and 700 K are $0.02\mathrm{s}^{-1}$ and $0.07\mathrm{s}^{-1}$, respectively. Determine the activation energy ($E_{\mathrm{a}}$) and the pre-exponential factor ($A$) for this reaction.
Solution:
$\log \frac {k _ {2}}{k _ {1}} = \frac {E _ {\mathrm {a}}}{2 . 3 0 3 R} \left[ \frac {T _ {2} - T _ {1}}{T _ {1} T _ {2}} \right]$
$\log \frac {0 . 0 7}{0 . 0 2} = \left(\frac {E _ {\mathrm {a}}}{2 . 3 0 3 \times 8 . 3 1 4 \mathrm {J K} ^ {- 1} \mathrm {m o l} ^ {- 1}}\right) \left[ \frac {7 0 0 - 5 0 0}{7 0 0 \times 5 0 0} \right]$
$0. 5 4 4 = E _ {\mathrm {a}} \times 5. 7 1 4 \times 1 0 ^ {- 4} / 1 9. 1 5$
$E _ {\mathrm {a}} = 0. 5 4 4 \times 1 9. 1 5 / 5. 7 1 4 \times 1 0 ^ {- 4} = 1 8 2 3 0. 8 \mathrm {J}$
Since
$k = A e ^ {- E a / R T}$
$0. 0 2 = A e ^ {- 1 8 2 3 0. 8 / 8. 3 1 4 \times 5 0 0}$
$A = 0. 0 2 / 0. 0 1 2 = 1. 6 1$
🧪 Example 3.10
Consider the decomposition of ethyl iodide, a first-order reaction represented by:
$\mathrm {C} _ {2} \mathrm {H} _ {5} \mathrm {I} (\mathrm {g}) \rightarrow \mathrm {C} _ {2} \mathrm {H} _ {4} (\mathrm {g}) + \mathrm {H I} (\mathrm {g})$
At 600 K, its rate constant is $1.60 \times 10^{-5} \mathrm{~s}^{-1}$, and its activation energy is 209 kJ/mol. Compute the reaction's rate constant when the temperature is 700 K.
Solution:
We know that
$\log k_2 - \log k_1 = \frac{E_a}{2.303R}\left(\frac{1}{T_1} - \frac{1}{T_2}\right)$
$\log k_2 = \log k_1 + \frac{E_a}{2.303R}\left(\frac{1}{T_1} - \frac{1}{T_2}\right)$
$\log k_2 = \log(1.60\times10^{-5}) + \frac{209000}{2.303\times8.314}\left(\frac{1}{600} - \frac{1}{700}\right)$
$\log k_2 = -4.796 + 2.599 = -2.197$
$k_2 = 6.36\times10^{-3}\ \mathrm{s^{-1}}$
3.4.1 Effect of Catalyst
A substance that accelerates the pace of a chemical reaction, yet remains chemically unaltered throughout the process, is defined as a catalyst. For instance, manganese dioxide ($\mathrm{MnO}_2$) significantly enhances the reaction rate of the following process.
$2 \mathrm {K C l O} _ {3} \xrightarrow {\mathrm {M n O} _ {2}} 2 \mathrm {K C L} + 3 \mathrm {O} _ {2}$
When an additive diminishes the reaction velocity, it is termed an inhibitor, and the designation "catalyst" is inappropriate. The mechanism by which a catalyst operates can be elucidated through the intermediate complex theory. This theoretical framework posits that a catalyst engages in the chemical process by establishing transient linkages with reactant species, thereby generating an intermediate complex. This complex is inherently unstable and ephemeral, subsequently breaking down to furnish the final products and regenerate the catalyst.
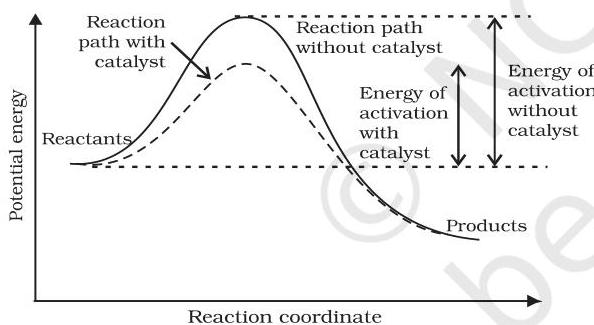
Fig. 3.11: Effect of catalyst on activation energy
The prevailing understanding is that catalysts facilitate an alternative reaction route or mechanistic sequence, which involves diminishing the activation energy separating the reactants from the products, consequently reducing the potential energy barrier, as visually represented in Fig. 3.11.
As indicated by the Arrhenius equation (3.18), a decrement in the activation energy value directly correlates with an augmented reaction rate.
A minute quantity of a catalytic agent possesses the capacity to process a substantial volume of reactants. A catalyst exerts no influence on the Gibbs free energy, $\Delta G$, of a reaction. Its action is limited to spontaneous reactions, and it does not facilitate non-spontaneous processes. Furthermore, observations reveal that a catalyst does not modify the equilibrium constant of a reaction; instead, its function is to accelerate the achievement of equilibrium. This occurs because it equally enhances the rates of both the forward and reverse reactions, ensuring that while the ultimate equilibrium state is unaltered, it is established more rapidly.
3.5 Collision Theory of Chemical Reactions
While the Arrhenius equation is applicable across a broad spectrum of conditions, the collision theory, formulated by Max Trautz and William Lewis between 1916 and 1918, offers a more profound understanding of the energetic and mechanistic facets of chemical reactions. This theory is grounded in the kinetic theory of gases. It posits that reactant molecules are
considered to be rigid spheres, and a reaction is hypothesized to occur when these molecules impact one another. The term "collision frequency
" (Z) refers to the number of collisions occurring per second within a unit volume of the reaction mixture. Activation energy, a concept previously discussed, represents another critical factor influencing the pace of chemical reactions. For an elementary bimolecular reaction expressed as:
$\mathrm {A} + \mathrm {B} \rightarrow \text {Products}$
the rate of reaction can be mathematically represented as:
$\operatorname {Rate} = Z _ {\mathrm {AB}} \mathrm {e} ^ {- E _ {\mathrm {a}} / R T} \tag {3.23}$
Here, $Z_{\mathrm{AB}}$ denotes the collision frequency between reactants A and B, while $\mathrm{e}^{-E_{\mathrm{a}} / RT}$ signifies the fraction of molecules possessing energy equal to or surpassing $E_{\mathrm{a}}$. A comparison of Equation (3.23) with the Arrhenius equation reveals a relationship between the pre-exponential factor A and collision frequency.
Equation (3.23) provides reasonably accurate predictions for the rate constants of reactions involving atomic species or straightforward molecules. However, for more intricate molecules, considerable discrepancies are observed. This divergence stems from the fact that not all collisions result in product formation. Collisions deemed "effective" are those where molecules collide with adequate kinetic energy (termed "threshold energy"*) and in the correct spatial orientation, thereby enabling the cleavage of existing bonds between reacting species and the subsequent formation of new bonds to yield products.
For instance, the synthesis of methanol from bromoethane is contingent on the orientation of the reactant molecules, as illustrated in Fig. 3.12. A proper alignment of reactant molecules facilitates bond formation, whereas an improper orientation merely causes them to rebound without producing any reaction.
To account for these effective collisions, an additional parameter, $P$, known as the probability or steric factor, is introduced. This factor incorporates the requirement that molecules must be suitably oriented during a collision. Thus, the rate equation becomes:
$\mathrm {Rate} = P Z _ {\mathrm {AB}} \mathrm {e} ^ {- E _ {\mathrm {a}} / R T}$
Consequently, according to collision theory, both the activation energy and the correct molecular orientation collectively establish the criteria for an effective collision and, by extension, dictate the rate of a chemical reaction.
Collision theory does possess certain limitations, notably its simplification of atoms and molecules as hard spheres, which overlooks their inherent structural characteristics. More comprehensive details regarding this theory and other theoretical frameworks will be explored in advanced academic courses.
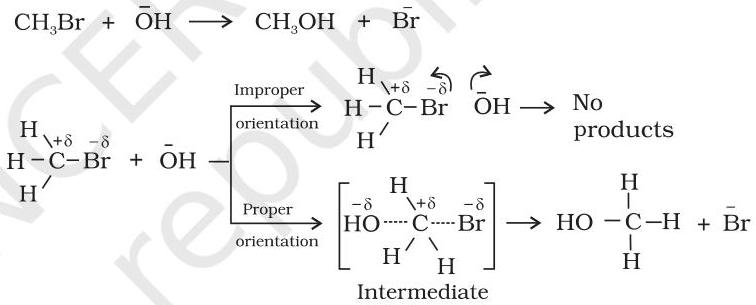
Fig. 3.12: Diagram showing molecules having proper and improper orientation
Intext Questions
3.7 What will be the effect of temperature on rate constant?
3.8 The rate of the chemical reaction doubles for an increase of 10K in absolute temperature from 298K. Calculate $E_{\mathrm{a}}$.
3.9 The activation energy for the reaction
$2 \mathrm{HI}(\mathrm{g}) \rightarrow \mathrm{H}_2 + \mathrm{I}_2(\mathrm{g})$
is 209.5 kJ mol$^{-1}$ at 581K. Calculate the fraction of molecules of reactants having energy equal to or greater than activation energy?
Summary
The discipline of chemical kinetics is dedicated to investigating chemical reactions with respect to their reaction rates, the influence of various parameters, the rearrangement of atomic configurations, and the formation of intermediate species. The rate of a reaction quantifies the decrease in reactant concentration or the increase in product concentration per unit time. This rate can be expressed as an instantaneous value at a particular moment or as an average value over an extended time interval. Numerous factors, such as temperature, reactant concentrations, and the presence of a catalyst, exert an influence on the reaction's velocity. The mathematical formulation describing a reaction's rate is known as the rate law, which must be empirically derived rather than theoretically foreseen. The order of a reaction with respect to a specific reactant is defined by the exponent of its concentration term within the rate law expression. The overall reaction order is the aggregate of these exponents for the concentration terms of all participating reactants. The rate constant serves as the proportionality coefficient within the rate law. Both the rate constant and the reaction order are ascertainable through either the rate law itself or its corresponding integrated form. Molecularity applies exclusively to elementary reactions, with its possible values restricted to integers between 1 and 3, contrasting with reaction order, which can be 0, 1, 2, 3, or even a non-integer. For an elementary reaction, the molecularity and the reaction order are numerically equivalent.
The relationship between temperature and rate constants is elucidated by the Arrhenius equation ($k = A\mathrm{e}^{-E_{\mathrm{a}}/RT}$). Here, $E_{\mathrm{a}}$ represents the activation energy, defined as the energy disparity between the transition state (activated complex) and the initial reactant species, while $A$ (known as the Arrhenius factor or pre-exponential factor) signifies the frequency of molecular collisions. This equation distinctly demonstrates that an elevation in temperature or a reduction in $E_{\mathrm{a}}$ will result in an accelerated reaction rate. Furthermore, the introduction of a catalyst diminishes the activation energy by furnishing an alternative reaction pathway. Collision theory introduces an additional parameter, $P$, termed the steric factor, which accounts for the spatial orientation of colliding molecules and its crucial role in facilitating effective collisions. This factor consequently modifies the Arrhenius equation to $k = PZ_{\mathrm{AB}}\mathrm{e}^{-E_{\mathrm{a}}/RT}$.
(i) $3\mathrm{NO}(\mathrm{g})\rightarrow \mathrm{N}_2\mathrm{O}$ (g) Rate $= k[\mathrm{NO}]^{2}$ (ii) $\mathrm{H}2\mathrm{O}2$ (aq) $+3\mathrm{I}^{-}$ (aq) $+2\mathrm{H}^{+}\rightarrow 2\mathrm{H}{2}\mathrm{O}$ (l) $+\mathrm{I}{3}^{-}$ Rate $= k[\mathrm{H}_2\mathrm{O}_2][\mathrm{I}^{-}]$ (iii) $\mathrm{CH}_3\mathrm{CHO}$ (g) $\rightarrow \mathrm{CH}_4$ (g) $+\mathrm{CO}(\mathrm{g})$ Rate $= k[\mathrm{CH}_3\mathrm{CHO}]^{3 / 2}$ (iv) $\mathrm{C}_2\mathrm{H}_5\mathrm{Cl}$ (g) $\rightarrow \mathrm{C}_2\mathrm{H}_4$ (g) $+\mathrm{HCl}$ (g) Rate $= k[\mathrm{C}_2\mathrm{H}_5\mathrm{Cl}]$
$2 \mathrm {A} + \mathrm {B} \rightarrow \mathrm {A} _ {2} \mathrm {B}$
The reaction rate is governed by the expression: rate $= k[\mathrm{A}][\mathrm{B}]^2$, where the rate constant $\mathrm{k}$ is $2.0 \times 10^{-6} , \mathrm{mol}^{-2} , \mathrm{L}^2 , \mathrm{s}^{-1}$. Determine the initial reaction rate when the concentrations are $[\mathrm{A}] = 0.1 , \mathrm{mol} , \mathrm{L}^{-1}$ and $[\mathrm{B}] = 0.2 , \mathrm{mol} , \mathrm{L}^{-1}$. Subsequently, calculate the reaction rate once the concentration of $[\mathrm{A}]$ has decreased to $0.06 , \mathrm{mol} , \mathrm{L}^{-1}$.
$\mathrm {R a t e} = k \left[ \mathrm {C H} _ {3} \mathrm {O C H} _ {3} \right] ^ {3 / 2}$
In a sealed container, the reaction rate can be monitored via the increase in pressure. Consequently, the rate may alternatively be articulated using the partial pressure of dimethyl ether, as shown:
$\mathrm {R a t e} = k \left(p _ {\mathrm {C H} _ {3} \mathrm {O C H} _ {3}}\right) ^ {3 / 2}$
Assuming pressure is quantified in bar and time in minutes, identify the appropriate units for the reaction rate and the rate constant.
(i) increased by a factor of two? (ii) decreased by half?
| t/s | 0 | 30 | 60 | 90 |
|---|---|---|---|---|
| [A]/ mol L$^{-1}$ | 0.55 | 0.31 | 0.17 | 0.085 |
Determine the average reaction rate within the time span ranging from 30 to 60 seconds.
(i) Formulate the differential rate law for this reaction. (ii) What consequence does tripling the concentration of B have on the reaction rate? (iii) How is the reaction rate altered if the concentrations of both A and B are simultaneously doubled?
| A/ mol L$^{-1}$ | 0.20 | 0.20 | 0.40 |
|---|---|---|---|
| B/ mol L$^{-1}$ | 0.30 | 0.10 | 0.05 |
| r₀/mol L$^{-1}$s$^{-1}$ | 5.07 × 10⁻⁵ | 5.07 × 10⁻⁵ | 1.43 × 10⁻⁴ |
What is the reaction order concerning reactant A and reactant B, respectively?
$2 \mathrm {A} + \mathrm {B} \rightarrow \mathrm {C} + \mathrm {D}$
| Experiment | [A]/mol L$^{-1}$ | [B]/mol L$^{-1}$ | Initial rate of formation of D/mol L$^{-1}$ min$^{-1}$ |
|---|---|---|---|
| I | 0.1 | 0.1 | 6.0 × 10⁻³ |
| II | 0.3 | 0.2 | 7.2 × 10⁻² |
| III | 0.3 | 0.4 | 2.88 × 10⁻¹ |
| IV | 0.4 | 0.1 | 2.40 × 10⁻² |
Determine the rate law and the rate constant for the reaction.
| Experiment | [A]/ mol L$^{-1}$ | [B]/ mol L$^{-1}$ | Initial rate/ mol L$^{-1}$ min$^{-1}$ |
|---|---|---|---|
| I | 0.1 | 0.1 | 2.0 × 10⁻² |
| II | - | 0.2 | 4.0 × 10⁻² |
| III | 0.4 | 0.4 | - |
| IV | - | 0.2 | 2.0 × 10⁻² |
(i) $200\mathrm{s}^{-1}$
(ii) $2\mathrm{min}^{-1}$
(iii) 4 years$^{-1}$
$\left[ 2 \mathrm {N} _ {2} \mathrm {O} _ {5} \rightarrow 4 \mathrm {N O} _ {2} + \mathrm {O} _ {2} \right]$
| t/s | 0 | 400 | 800 | 1200 | 1600 | 2000 | 2400 | 2800 | 3200 |
|---|---|---|---|---|---|---|---|---|---|
| 10$^2$ × [N₂O₅]/mol L$^{-1}$ | 1.63 | 1.36 | 1.14 | 0.93 | 0.78 | 0.64 | 0.53 | 0.43 | 0.35 |
(i) Graph the concentration of $\left[\mathrm{N}{2} \mathrm{O}{5}\right]$ as a function of time.
(ii) Determine the reaction's half-life period.
(iii) Construct a plot of $\log[\mathrm{N}_2\mathrm{O}_5]$ versus time.
(iv) Formulate the rate law expression.
(v) Compute the value of the rate constant. (vi) Determine the half-life period using the calculated rate constant $k$, and compare this value with the result from part (ii).
| t (sec) | P(mm of Hg) |
|---|---|
| 0 | 35.0 |
| 360 | 54.0 |
| 720 | 63.0 |
Determine the rate constant for this process.
$\mathrm{SO}_2\mathrm{Cl}_2(\mathrm{g}) \rightarrow \mathrm{SO}_2(\mathrm{g}) + \mathrm{Cl}_2(\mathrm{g})$
| Experiment | Time/s⁻¹ | Total pressure/atm |
|---|---|---|
| 1 | 0 | 0.5 |
| 2 | 100 | 0.6 |
Determine the reaction rate when the total pressure reaches 0.65 atm.
| T/°C | 0 | 20 | 40 | 60 | 80 |
|---|---|---|---|---|---|
| 10$^5$ × k/s$^{-1}$ | 0.0787 | 1.70 | 25.7 | 178 | 2140 |
Construct a plot of $\ln k$ against $1/T$ to determine the pre-exponential factor ($A$) and the activation energy ($E_{\mathrm{a}}$). Subsequently, estimate the rate constants at $30^{\circ}\mathrm{C}$ and $50^{\circ}\mathrm{C}$.
$k = (4.5 \times 10^{11} \mathrm{~s}^{-1}) e^{-28000K/T}$
Determine the value of the activation energy, $E_{\mathrm{a}}$.
Chemical Kinetics
$\log k = 14.34 - 1.25 \times 10^4 , \mathrm{K}/T$
Compute the activation energy ($E_{\mathrm{a}}$) for this reaction. Furthermore, identify the temperature at which its half-life will be $256$ minutes.
{'number': '3.1', 'text': 'From the rate expression for the following reactions, determine their order of reaction and the dimensions of the rate constants.'}
{'number': '3.2', 'text': 'Consider the chemical transformation:'}
{'number': '3.3', 'text': 'The catalytic breakdown of $\mathrm{NH}_3$ occurring on a platinum surface is characterized as a zero-order reaction. Given that the rate constant $k = 2.5\times 10^{-4}\mathrm{mol}^{-1}\mathrm{L}\mathrm{s}^{-1}$, determine the rates at which $\mathbf{N}2$ and $\mathrm{H}{2}$ are generated.'}
{'number': '3.4', 'text': "Dimethyl ether undergoes decomposition, yielding $\mathrm{CH}_4$, $\mathrm{H}_2$, and CO. The reaction's rate is defined by the expression:"}
{'number': '3.5', 'text': 'Enumerate the variables that influence the speed of a chemical process.'}
{'number': '3.6', 'text': 'If a chemical reaction exhibits second-order kinetics concerning a specific reactant, how would the reaction rate be impacted if the concentration of that reactant is:'}
{'number': '3.7', 'text': "Describe the influence of temperature on a reaction's rate constant. Furthermore, how can this temperature-dependent relationship for the rate constant be mathematically expressed?"}
{'number': '3.8', 'text': 'For a pseudo first-order reaction conducted in an aqueous medium, the experimental data presented below were acquired:'}
{'number': '3.9', 'text': 'Consider a reaction that is first order with respect to reactant A and second order with respect to reactant B.'}
{'number': '3.10', 'text': 'For a reaction involving reactants A and B, the initial reaction rate $(\mathbf{r}_0)$ was experimentally determined under various initial concentrations of A and B, as detailed in the table below:'}
{'number': '3.11', 'text': 'The subsequent experimental data were gathered during investigations into the kinetics of the reaction:'}
{'number': '3.12', 'text': 'For a reaction involving species A and B, the kinetics are first-order with respect to A and zero-order with respect to B. Complete the missing entries in the table provided below:'}
{'number': '3.13', 'text': 'Compute the half-life period for a first-order reaction using the rate constants provided below:'}
{'number': '3.14', 'text': 'The radioactive decay of $^{14}\mathrm{C}$ exhibits a half-life of 5730 years. An archaeological artifact composed of wood was found to possess $80\%$ of the $^{14}\mathrm{C}$ concentration typically found in a living tree. Estimate the age of the sample.'}
{'number': '3.15', 'text': 'The following experimental observations pertain to the gas-phase decomposition of $\mathrm{N}_2\mathrm{O}5$ at 318K:'}
{'number': '3.16', 'text': 'A first-order reaction possesses a rate constant of $60~\mathrm{s}^{-1}$. Calculate the duration required for the initial reactant concentration to diminish to $1/16^{\text{th}}$ of its original value.'}
{'number': '3.17', 'text': 'A significant product generated during a nuclear detonation is $^{90}\mathrm{Sr}$, characterized by a half-life of 28.1 years. If $1\mu\mathrm{g}$ of $^{90}\mathrm{Sr}$ were to be incorporated into the skeletal structure of a neonate, replacing calcium, determine the residual quantity after durations of 10 years and 60 years, assuming no metabolic excretion.'}
{'number': '3.18', 'text': 'For a first-order reaction, demonstrate that the time necessary for $99\%$ completion is precisely double the time required for $90\%$ completion.'}
{'number': '3.19', 'text': 'A first-order reaction undergoes $30\%$ decomposition over a period of $40\mathrm{min}$. Determine its half-life, $t{1/2}$.'}
{'number': '3.20', 'text': 'Empirical data for the decomposition of azoisopropane, producing hexane and nitrogen at 543 K, are presented below:'}
{'number': '3.21', 'text': 'The ensuing data were collected for the first-order thermal decomposition of $\mathrm{SO}_2\mathrm{Cl}_2$ under conditions of constant volume:'}
{'number': '3.22', 'text': 'The decomposition of $\mathrm{N}_2\mathrm{O}5$ exhibits the following rate constant values across a range of temperatures:'}
{'number': '3.23', 'text': 'For the breakdown of hydrocarbons, the rate constant is determined to be $2.418 \times 10^{-5} \mathrm{s}^{-1}$ at a temperature of $546 \mathrm{K}$. Given an activation energy of $179.9 \mathrm{kJ} / \mathrm{mol}$, calculate the magnitude of the pre-exponential factor.'}
{'number': '3.24', 'text': 'For a specific reaction, $\mathrm{A} \rightarrow$ Products, the rate constant $k$ is $2.0 \times 10^{-2} \mathrm{s}^{-1}$. Determine the concentration of reactant $\mathrm{A}$ that persists after $100 \mathrm{s}$, assuming an initial concentration of $1.0 \mathrm{mol} \mathrm{~L}^{-1}$ for $\mathrm{A}$.'}
{'number': '3.25', 'text': 'In an acidic medium, sucrose undergoes decomposition into glucose and fructose, following a first-order kinetic pathway with a half-life ($t{1/2}$) of $3.00$ hours. Ascertain the fraction of the initial sucrose sample that will be left after a duration of $8$ hours.'}
{'number': '3.26', 'text': 'The kinetic behavior for the decomposition of a hydrocarbon is described by the equation:'}
{'number': '3.27', 'text': 'The rate constant for the first-order decomposition of hydrogen peroxide ($\mathrm{H}_2\mathrm{O}2$) is expressed by the subsequent equation:'}
{'number': '3.28', 'text': 'For the decomposition of reactant $\mathrm{A}$ into products, the rate constant $k$ is $4.5 \times 10^{3} \, \mathrm{s}^{-1}$ at $10^{\circ}\mathrm{C}$, with an associated activation energy of $60 \, \mathrm{kJ} \, \mathrm{mol}^{-1}$. Determine the temperature at which the rate constant $k$ would attain a value of $1.5 \times 10^{4} \, \mathrm{s}^{-1}$.'}
{'number': '3.29', 'text': 'For a first-order reaction, the duration needed for $10\%$ completion at $298 \mathrm{K}$ is equivalent to the duration required for $25\%$ completion at $308 \mathrm{K}$. Given that the pre-exponential factor ($A$) is $4 \times 10^{10} \, \mathrm{s}^{-1}$, compute the rate constant ($k$) at $318 \mathrm{~K}$ and the activation energy ($E{\mathrm{a}}$).'}
{'number': '3.30', 'text': "A reaction's rate increases by a factor of four when the temperature is elevated from $293 \mathrm{K}$ to $313 \mathrm{K}$. Determine the activation energy of this reaction, assuming its value remains constant irrespective of temperature fluctuations."}
Answers to Some Intext Questions
3.1 $r_{\mathrm{av}} = 6.66 \times 10^{-6} , \mathrm{Ms}^{-1}$
3.2 Rate of reaction = rate of disappearance of A
$= 0.005 , \mathrm{mol} , \mathrm{litre}^{-1} , \mathrm{min}^{-1}$
3.3 Order of the reaction is 2.5
3.4 $X \rightarrow Y$
$\text{Rate} = k[X]^2$
The rate will increase 9 times
3.5 $t = 444 , \mathrm{s}$
3.6 $1.925 \times 10^{-4} , \mathrm{s}^{-1}$
3.8 $E_{\mathrm{a}} = 52.897 , \mathrm{kJ} , \mathrm{mol}^{-1}$
3.9 $1.471 \times 10^{-19}$